This Message Is From an Untrusted Sender
You have not previously corresponded with this sender.
Hello,
I am working on an integrated model utilizing a ~3.4-angstrom cryo-EM structure and a crosslink dataset. Comparing the structure produced by my 400-gaussian gmm.txt file to my cryo map mrc, I am concerned that many details are lost in translation. Are there
ways to improve the precision of the gmm.txt file?
For instance, would any of the following tools from the usage guide be helpful, and how would I use them? [-t {spherical,tied,diag,full}] [-m OUT_MAP] [-a APIX] [-n NUM_SAMPLES] [-i NUM_ITER] [-s THRESHOLD] [-f FORCE_RADII] [-w FORCE_WEIGHT] [-e] [-d] [-k]
[-x CHAIN] [-z]
[-x CHAIN] [-z]
This is the command I am currently using in IMP 2.24:
python create_gmm.py \
cluster01.mrc \
400 \
cluster01_400.gmm.txt
Pictures left to right of my spherical, 1-MDa complex in mrc and gmm.mrc:
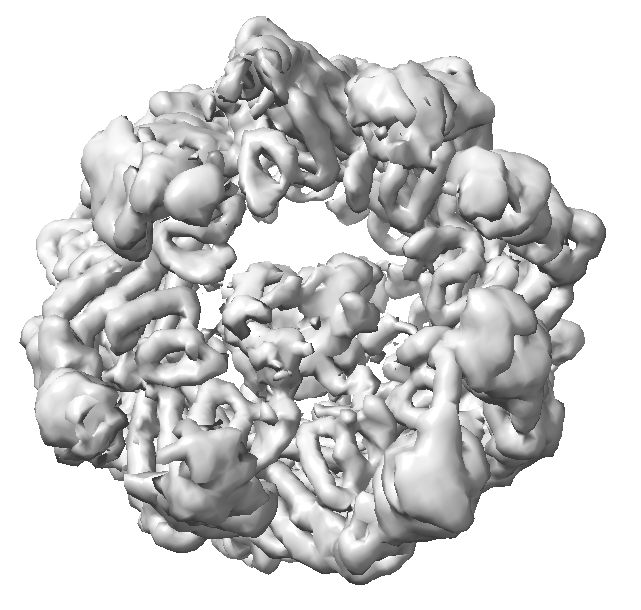
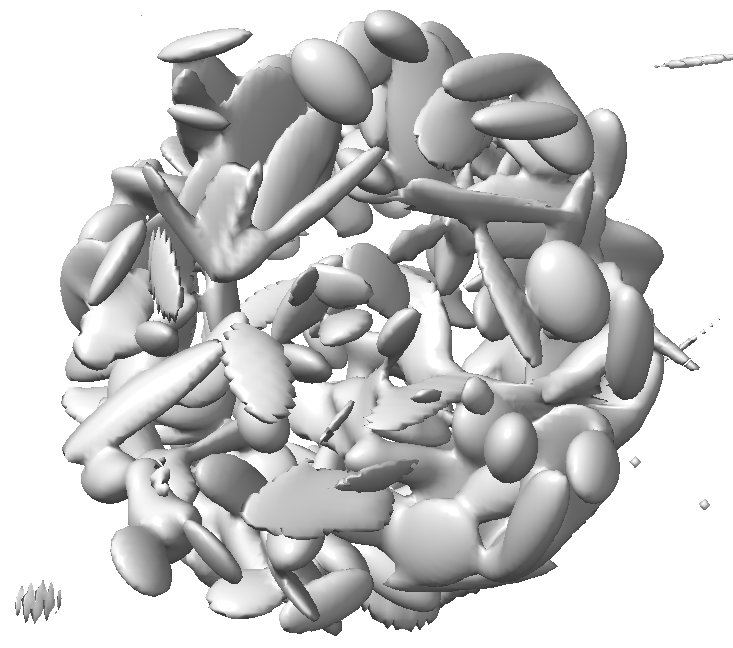
I did consult AI and received this recommended improvement. Would you add any suggestions to this?
python create_gmm.py \
-s <chimerax_viewing_threshold> \
-n 4000000 \
-i 300 \
-t full \
-m cluster01_400.gmm.mrc \
-a <map_resolution_in_angstroms> \
cluster01.mrc 400 cluster01_400.gmm.txt
Thank you so much for your help,
Cecilia Sanders
6th year Ph.D. student, Barry Willardson Lab
BYU Dept. of Chemistry and Biochemistry
IMP-users mailing list; unsubscribe